【Q&A】| 瑞士製藥監督機構關於EU GMP附錄1的問答(上篇)
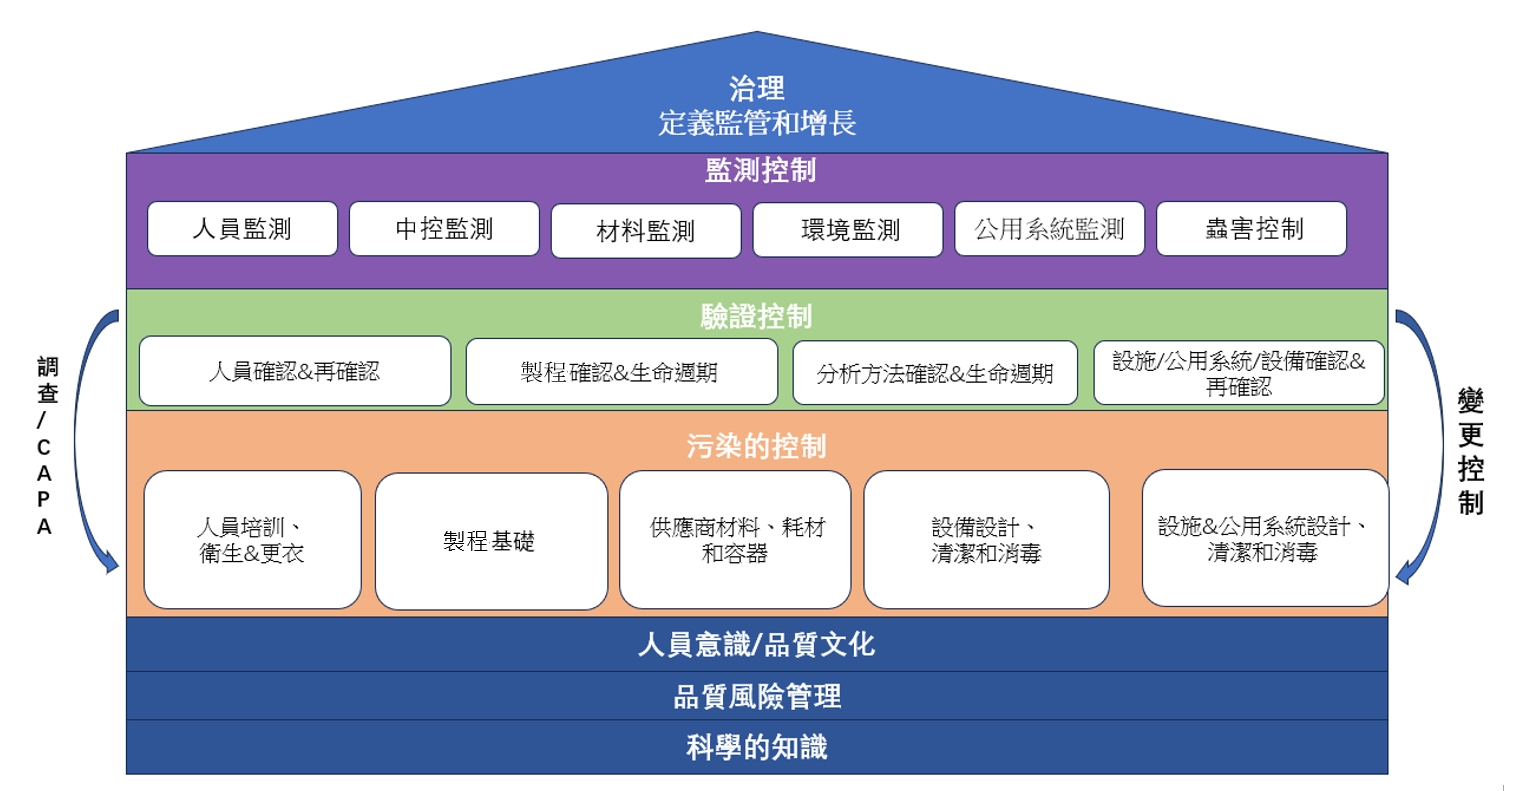
瑞士製藥監督機構針對EU GMP Annex1的解讀問答主要包含其中7章內容,涵蓋範圍Scope、廠房Premises、公用設施Utilities、人員/訓練Personnel/Training、生產與特定技術Production and Specific Technologies、環境監測與製程監測Environmental & Process monitoring以及品質管制Quality Control (QC),本期主要說明其中前四項內容,內容如下:
Interpretation: Questions and Answers
解讀:問題與解答
4.1 Scope(Annex l,chapter 1)
範圍(附錄 1,第1章)
Q1: Does Annex 1 apply without restriction also to ATMPs or are only defined aspects of Annex 1 to be followed for some specific product types, such as for example the allogeneic and autologous cell therapy products?
附錄1是否也不受限制地適用於 ATMP,還是僅針對某些特定產品類型(例如異體和自體細胞治療產品)才遵循附錄 1 的規定方面?
A: It is recognised that ATMPs cover a very heterogeneous range of products and that for some of these products, due to their nature and manufacturing technology, specific considerations are required. This certainly applies, for example, to the allogeneic and autologous cell therapy pro-ducts, which are to be manufactured under conditions suitable to avoid microbial contaminations, but which usually cannot be terminally sterilised or sterile filtered. In addition, such products are made from unsterile patient material. Specifically with regard to cellular therapy, Annex 2A, para-graph 5.29(b), requires that aseptic processing be maintained from the time of procurement of cells through manufacturing and administration back into the patient. Annex 2A refers to Annex 1 several times (e.g., in connection with the requirements for the provision of systems for closed processing), but implies the possibility of exceptions from applying the requirements of Annex 1. It must also be taken into account that Annex 2A became valid in May 2021, i.e., more than one year before the publication of the new Annex 1 version.
答:一般認為,ATMP 涵蓋的產品種類相當多元,其中部分產品因其性質與製造技術,需要特別考量。這當然適用於異體與自體細胞治療產品;這些產品應在適當條件下製造,以避免微生物污染,但通常無法進行最終滅菌或除菌過濾。此外,這類產品是以非無菌的病患材料製成。就細胞治療而言,附錄 2A 第 5.29(b) 段要求,從細胞取得、製造到再施用回病患體內,皆必須維持無菌操作。附錄 2A 多次提及附錄 1(例如關於提供封閉式處理系統的要求),但也暗示可能有不適用附錄 1 要求的例外情形。另須考量的是,附錄 2A 於 2021 年 5 月生效,早於新版附錄 1 發布一年以上。
It is expected that ATMP manufacturers, based on the knowledge of their manufacturing processes and the execution of detailed risk analyses covering all process steps, materials and systems, develop and implement Contamination Control Strategies suitable to avoid or largely minimise risks of product contaminations. Justification must be given for any exceptions to the requirements of Annex 1.
ATMP 製造商應依據其對製造製程的瞭解,以及涵蓋所有製程步驟、原料與系統的詳細風險分析,制定並執行適當的污染管制策略,以避免或盡可能降低產品污染風險。對於任何未適用附錄 1 要求的例外情形,均必須提出理由說明。
4.2 Premises (Annex 1, Chapter 4)
廠房(附錄 1,第 4 章)
Q2: Are grade A and B cleanrooms required to have separate air-locks for material and personnel and must the flows in such air-locks be strictly unidirectional?
A 級和 B 級潔淨室是否需要為物料與人員分別設置氣鎖,且氣鎖中的人員與物料動線是否必須嚴格為單向流?
A: In general, it is expected that new facilities have for grade A and B zones segregated airlocks for personnel and material and that the flows in such airlocks are unidirectional (i.e., separate MALs for transport into and out of the cleanroom and separate PALs for personnel entry and exit). Existing facilities that do not have such airlock separation must ensure that, as a minimum, temporary separation of the flows in the airlocks is guaranteed and that the situation is covered by scientifically sound risk analysis also assessing the need of additional technical or organizational measures. The rationale for not applying physical separation of the above-mentioned flows through segregated airlocks and the risk assessment on which it is based must be integrated in the overall contamination control strategy.
答:一般而言,新建設施的 A 級和 B 級區域,預期應分別設置人員與物料氣鎖,且氣鎖中的動線應為單向(亦即物料進出潔淨室應分別使用獨立的物料氣鎖,人員進出潔淨室應分別使用獨立的人員氣鎖)。未設置此類氣鎖分流的既有設施,至少必須確保氣鎖內的人員與物料動線可暫時分隔,並應以科學合理的風險分析涵蓋此情況,同時評估是否需要額外的技術或組織措施。未透過分隔氣鎖對上述動線進行實體分隔的理由,以及其所依據的風險評估,必須納入整體污染管制策略。
Q3:Paragraph 4.12, point ii, states that only materials and equipment that are on an approved list and that have been assessed during validations of the transfer process, should be transferred into grade A or grade B areas via airlocks or pass-through hatches. What does “validation of the transfer process” mean, for example for the material transfer into an isolator?
4.12第 ii 項規定,只有列入核准清單,並於轉移流程確效期間完成評估的物料與設備,才可透過氣鎖或傳遞窗轉移至 A 級或 B 級區域。例如,對於將物料轉移至隔離器而言,「轉移流程確效」是什麼意思?
A: As mentioned in paragraph 4.10 of Annex 1, the transfer of materials, equipment or components into and out of a cleanroom (incl. the critical zone within a grade A environment), represents one of the greatest potential sources and risks of contamination.
答:正如附錄 1 的 4.10 所述,材料、設備或部件進出潔淨室(包括 A 級環境內的關鍵區域)的轉移是最大的潛在污染源和風險之一。
In order to minimise such risks, great care must be taken in particular when defining the technical and procedural measures associated with the transfer of materials/equipment into an aseptic processing area. Only in relatively rare cases it is possible to bring materials into an isolator before it is sealed and bio-decontaminate them together with the isolator using a validated VHP treatment (only possible for small batches and if materials are resistant to VHP). In the majority of cases, however, it is necessary to transfer materials to an isolator that has already been decontaminated. For this, all materials must first be sterilised and then moved through the physical barrier of the isolator in such a way that the sterility of the goods and the integrity of the isolator are maintained. Regardless of the technology used (e.g., usage of double-door sterilisers upstream of the isolator, use of transfer isolators or of rapid transfer port technology), the entire transfer process must be considered within detailed risk analysis and be part of the overall contamination control strategy. In addition, appropriate control mechanisms must be defined to monitor the maintenance of the integrity and functionality of the systems (e.g., measurement of differential pressure and control of door interlocks between adjacent zones of the transfer system). The technical solutions must be covered by appropriate equipment/system qualifications (incl. smoke studies if applicable) and sterilisation validations and the suitability of the entire transfer process must be verified through APS (validations and also regular APS). Appropriate qualification measures and APS must also be used to demonstrate that the egress of materials from the isolator does not affect the maintenance of the grade A zone requirements.
為盡可能降低此類風險,在界定與物料/設備轉移至無菌操作區相關的技術與程序性措施時,必須特別謹慎。僅在相對少見的情況下,才可能於隔離器密封前將物料置入隔離器內,並使用經確效的 VHP 處理,將其與隔離器一併進行生物去污(僅適用於小批量且物料可耐受 VHP 的情況)。然而,多數情況下,必須將物料轉移至已完成去污的隔離器。為此,所有物料必須先經滅菌,再透過隔離器的實體屏障移入,同時維持物品無菌性與隔離器完整性。無論採用何種技術(例如在隔離器上游使用雙門滅菌器、使用轉移隔離器或快速轉移介面技術),整個轉移流程均必須納入詳細風險分析,並成為整體污染管制策略的一部分。此外,必須界定適當的管制機制,以監測系統完整性與功能性的維持情形(例如壓差量測,以及轉移系統相鄰區域之間的門互鎖控制)。技術方案必須涵蓋適當的設備/系統驗證(如適用,包含煙流型態研究)與滅菌確效,並須透過 APS 確認整個轉移流程的適用性(確效及定期 APS)。亦必須採用適當的驗證措施與 APS,證明物料自隔離器移出不會影響 A 級區域要求的維持。
The arrangement of the installations, the processes carried out in it and the material movements must also be considered when defining the points to be sampled during PQ activities or during routine or event based environmental monitoring.
在PQ 活動期間,或於例行性或事件導向環境監測期間界定取樣點時,亦須考量設施配置、其中執行的製程,以及物料移動情形。
Q4: Is it always required to strictly adhere to the area cleanliness cascade (i.e., respecting the sequential order of cleanroom classes) for material transfer through airlocks or pass-through hatches or is it possible to skip a grade (e.g., moving from CNC directly to class C) under certain circumstances?
透過氣鎖或傳遞窗進行物料轉移時,是否一律必須嚴格遵循區域潔淨度級序(亦即遵守潔淨室等級的先後順序),或在特定情況下是否可跳級(例如從 CNC 直接轉入 C 級)?
A :Compliance with the cleanroom sequence for the transfer of materials via airlocks or pass-through hatches is expected to be fulfilled for zones A and B (exceptions from this rule are possible for sterility test rooms). For cleanroom areas with lower classification, it is principally feasible for materials to be transferred from one low zone (CNC) through an airlock or pass-through hatch directly into an area with two grades higher classification (grade C area), provided that suitable technical and/or procedural measures are established ensuring fulfilment of the cleanroom specifications in the respective areas. The adequacy of the established systems/procedures needs to be demonstrated by appropriate qualification activities and the results of regular environmental monitoring. The defined measures and the risk analyses on which they are based must be part of the CCS.
答:對於A 級和 B級區域(無菌試驗室可例外),透過氣鎖或傳遞窗轉移物料時,預期應符合潔淨室級序。對於較低等級的潔淨室區域,只要已建立適當的技術及/或程序性措施,確保各相關區域符合潔淨室規格,原則上可將物料從低等級區域(CNC)經由氣鎖或傳遞窗,直接轉移至高兩級的區域(C 級區域)。既有系統/程序的適當性,須透過適當的確認活動與定期環境監測結果加以證明。所界定的措施及其所依據的風險分析,必須納入污染管制策略(CCS)。
Q5: What are the expectations for older barrier technology systems that do not meet all the requirements according to the new Annex 1? By when do they have to be replaced or upgraded?
對於不符合新附錄 1 所有要求的舊屏障技術系統有何期望?到什麼時候必須更換或升級?
A: The company has to perform an in-depth internal evaluation of the current barrier technology and assess whether the installation, its cleanroom background and all related systems/procedures meet the requirements of the new Annex 1 or whether technical measures are required. If necessary, a project has to be initiated for example to upgrade the cleanroom used as background and install additional airlocks. From August 25th 2023, all barrier technology equipment not complying with the revised Annex 1 are considered deficient and deviations will be issued upon findings during inspections. Depending on the CAPA plan and interim risk reducing measures defined, an additional implementation timeline of approx. one year may be acceptable.
答:公司必須對現行屏障技術進行深入的內部評估,並評估該設施、其潔淨室背景環境及所有相關系統/程序是否符合新版附錄 1 要求,或是否需要採取技術措施。如有必要,必須啟動專案,例如升級作為背景環境的潔淨室並增設氣鎖。自 2023 年 8 月 25 日起,所有不符合修訂後附錄 1 要求的屏障技術設備,均視為存在缺失;檢查時如發現此類情形,將開立缺失。依據所訂定的 CAPA 計畫及暫行風險降低措施,額外約一年的執行時程可能可被接受。
Q6: What is meant in paragraph 4.20 by the need to take into consideration, among others, the “extent of automation” when carrying out CCS related risk assessments of an isolator?
第 4.20 段中,在對隔離器進行 CCS 相關風險評估時,需要考量「自動化程度」,其意義為何?
A :This refers to the inclusion in such CCS risk assessments of an evaluation of all automated functionalities and processes associated with the use of the isolator and the activities taking place in it (from cleaning and disinfection of the equipment, to the transport of materials into the isolator, their handling and the product filling, to the capping and removal of the filled containers). The use of a well-designed, automated, recipe-controlled and possibly robotised system, equipped with appropriate control and alarm systems, can increase the reproducibility of the operations and minimise both errors and manual interventions. Ideally, such risk analyses should al-ready be carried out as part of the design or se-lection of the isolator system and should be revised or supplemented during the lifecycle of the equipment, as knowledge and experience in-creases or in the event of changes to be implemented.
答:這是指在此類 CCS 風險評估中,納入對隔離器使用及其內部作業相關之所有自動化功能與流程的評估(從設備清潔與消毒、物料轉運至隔離器內、物料處理與產品充填,到已充填容器的軋蓋與移出)。使用設計良好、自動化、配方控制且可能採用機器人化的系統,並配備適當的控制與警報系統,可提升作業再現性,並盡可能降低錯誤與人工介入。理想情況下,此類風險分析應已作為隔離器系統設計或選型的一部分執行,並應於設備生命週期中,隨著知識與經驗增加,或在需要實施變更時進行修訂或補充。
Q7: Are manual operations accepted for bio-decontamination?
是否接受以手動操作進行生物去污?
A:The decontamination process for an isolator should always be an automatic process. How-ever, paragraph 4.22 refers to both RABS and isolators, and manual decontamination processes are most commonly encountered for RABS. Such manual processes must be de-signed in such a way to be reproducible and to cover the entire surface area of the equipment, and their robustness and effectiveness must be demonstrated by appropriate validation and by regular monitoring.
答:隔離器的去污程序原則上應為自動化程序。然而,4.22 同時提及 RABS與隔離器,而手動去污程序最常見於 RABS。此類手動程序的設計必須具備再現性,並涵蓋設備整個表面區域;其穩健性與有效性必須透過適當確效及定期監測加以證明。
Q8: “Evidence should also be available to demonstrate that the cleaning and bio-decontamination agents used do not have ad-verse impact on the product produced within the RABS or isolator”. How should this be demonstrated?
“亦應備有證據,證明所使用的清潔劑與生物去污劑,不會對 RABS 或隔離器內生產的產品造成不良影響”。應如何證明?
A : The cleaning or bio-decontamination procedure should include steps designed to effectively re-move cleaning agent or disinfectant (including sporicidal agent) residues from direct and indirect product contact surfaces within the RABS/isolator. The effectiveness of these steps should be demonstrated based on validation data. For isolators, validation data should be available to demonstrate that the residual amount of sporicidal agent is below the concentration that could be detrimental to the product quality and stability at the end of the sporicidal cycle. Control mechanisms must be defined to ensure that the conditions prevailing during validation are also maintained during routine production (e.g., compliance with the time after completion of the sporicidal cycle, resp. measurement of the peroxide concentration).
答:清潔或生物去污程序應包括能有效去除 RABS/隔離器內直接與間接產品接觸表面之清潔劑或消毒劑(包括殺孢子劑)殘留的步驟。這些步驟的有效性應依據確效資料證明。對於隔離器,應備有確效資料,證明在殺孢子循環結束時,殺孢子劑殘留量低於可能危害產品品質與安定性的濃度。必須界定管制機制,以確保例行生產期間亦能維持確效期間的條件(例如遵守殺孢子循環完成後的等待時間,或量測過氧化物濃度)。
The extent to which samples of the surfaces are to be taken during validation and analysed for disinfectant/ cleaning agent residues can be de-fined based on a risk assessment, taking into ac-count the risk of transferring such residues to the product or a product contacting surface of a packaging component.
確效期間是否需對表面進行取樣並分析消毒劑/清潔劑殘留,以及其取樣程度,可依據風險評估界定;評估時應考量此類殘留轉移至產品,或轉移至包材產品接觸表面的風險。
Q9: Is it acceptable that for barrier technology systems with unidirectional air flow other air speed and speed measurement positions are defined than those mentioned in Annex 1?
對於採用單向氣流的屏障技術系統,是否可界定不同於附錄 1 所述的其他風速範圍與風速量測位置?
A : The most important requirement for barrier technology systems stated in paragraph 4.30 is that the air velocity in unidirectional airflow systems must be defined in such a way that unidirectional and uniform airflow conditions prevail at the working positions where high-risk operations take place, suitable to protect the product and open components (e.g., containers) from contamination.
答:4.30 所述之屏障技術系統最重要的要求,是單向氣流系統中的風速必須加以界定,使執行高風險作業的工作位置能維持單向且均勻的氣流條件,以保護產品與開放式組件(如容器)免受污染。
The air speed range of 0.36 – 0.54 m/s is, as stated in the above paragraph itself, merely a guideline value that has been encountered in the pharmaceutical industry for decades.
如上述段落所述,0.36 – 0.54 m/s 的風速範圍僅為製藥產業多年來常見的指引值。
Annex 1, however, clearly allows for the establishment of alternative air speed ranges or measurements at different heights in the system than the working position, provided this is “scientifically justified in the CCS”. It is important that the suitability of the defined airflow conditions is proven by airflow visualisation studies (part of the system qualification) covering the entire sys-tem and that these are correlated with the respective defined air speed range at specified height/position. The air speed must be measured continuously during operations and kept within this defined range.
然而,附錄 1 明確允許在系統中不同於工作位置的高度建立替代風速範圍或量測方式,前提是其已「在 CCS 中提出科學依據」。重要的是,所界定氣流條件的適用性,須透過涵蓋整個系統的氣流可視化研究(屬於系統確認的一部分)加以證明,且這些研究應與指定高度/位置所界定的相應風速範圍建立關聯。作業期間必須連續量測風速,並將其維持於該界定範圍內。
4.3 Utilities (Annex 1, Chapter 6)
公用設施(附錄 1,第 6 章)
Q10: Water generation – reverse osmosis system: What are the requirements regarding the sanitization (disinfection) of the system?
製水:系統的消毒處理有何要求?
A: Paragraph 6.12 gives detailed guidance on the requirements. It is important that the system is designed to allow for routine sanitization / disinfection and a procedure is in place defining this regular preventive sanitization or disinfection of the RO-system. It should also include a regular change of membranes. The frequency of sanitization should be determined based on quality risk management principles and on the data gathered during the qualification of the system, and it should be reviewed at least annually taking into consideration the routine monitoring data.
答: 6.12 提供了有關要求的詳細指引。重要的是,系統設計應允許進行例行衛生化處理/消毒,並應建立程序,界定逆滲透系統的定期預防性衛生化處理或消毒。程序亦應包括定期更換濾膜。衛生化處理頻率應依據品質風險管理原則,以及系統確認期間所蒐集的資料決定,並應至少每年檢討一次,同時納入例行監測資料考量。
The system must continuously be maintained meaning that the sanitization also has be per-formed when no production is running or when no water is used for production.
系統必須持續維護,這意味著在不生產或生產不用水時也必須進行消毒。
Q11: What are the sampling requirements for regular ongoing monitoring of Water for Injection?
對注射用水定期持續監測的取樣要求是什麼?
A :A suitable sampling schedule should be in place to ensure that representative water samples are obtained for analysis on a regular basis. For WFI distribution system sampling plans are more important because microbial control must be much more stringent. In general, water sampling for microbial and bacterial endotoxin testing is expected to occur daily somewhere in the system, with each outlet being sampled periodically to characterize the quality of the water. The use of cold loops requires a much closer microbiological monitoring and special sanitization measures.
答:應建立適當的取樣計畫,以確保定期取得具代表性的水樣進行分析。對於 WFI 分配系統而言,取樣計畫更為重要,因為微生物管制必須更加嚴格。一般而言,系統中某處應每日進行微生物與細菌內毒素檢測用水樣取樣,並定期對每個使用點取樣,以確認水質特性。使用冷迴路系統時,需要更嚴密的微生物監測與特殊的衛生化處理措施。
Quality control sampling locations in the main distribution system should include all POUs, having also process control sampling be located before the first and after the last POU and at other specified worst-case locations. POU sampling plans should rotate through all use points on the system, with the expectation that samples are collected on a daily basis from various use points, and that all use points are sampled on a rotational basis. The loop return should be sampled each day of use of the system in order to provide additional assurance of the quality of water utilized in the manufacturing processes. For WFI, it is an expectation that water samples should be taken daily from a minimum of one POU, with all point of use tested weekly during the qualification phase. The final phase of qualification may form the basis for the ongoing sampling frequencies with the goal of ensuring that the system is maintained in a validated state. However, it has become good industry practice to continue to utilize the same sampling frequency beyond the completion of the performance qualification to collect sufficient historical data in order to justify adjusting the sampling frequency. The use of risk analysis tools coupled with stringent periodic data review may be used to alter the frequency of sampling. Any de-crease of the sampling frequency for routine monitoring should be based on historical data and should only occur when a large number of historical data is available to allow statistical analysis. Based on the outcome of analysis of data and on the regular review of the performance of the point of use or the system, and if operational SOPs are in place which ensure also an increase of the sampling frequency if indicated and regular maintenance activities, e.g. for all outlets, a less frequent sampling can be justified. The risk assessment should consider the fact that decreased sampling frequencies also results in a higher number of batches that will be put at risk and a problem may have a serious impact on supply of products for patients.
主要分配系統中的品質管制取樣位置應包括所有使用點,同時製程管制取樣點應設於第一個使用點之前、最後一個使用點之後,以及其他指定的最不利情境位置。使用點取樣計畫應輪流涵蓋系統中的所有使用點,預期每日從不同使用點採集樣品,並以輪替方式完成所有使用點取樣。系統使用期間,應每日對迴路回流進行取樣,以進一步確保製造製程中所使用水的品質。對於 WFI 而言,預期每日應至少自一個使用點採集水樣,並於確認階段每週檢測所有使用點。確認的最後階段可作為持續取樣頻率的依據,目標是確保系統維持於已確效狀態。然而,在性能確認完成後持續採用相同取樣頻率,以蒐集足夠歷史資料作為調整取樣頻率的合理依據,已成為業界良好作法。可使用風險分析工具並搭配嚴格的定期資料審查,以調整取樣頻率。任何例行監測取樣頻率的降低,均應依據歷史資料,且僅在具備大量歷史資料足以進行統計分析時方可執行。依據資料分析結果,以及對使用點或系統性能的定期審查,若已建立操作 SOP,並能確保在有需要時提高取樣頻率及執行定期維護活動(例如針對所有出水口),則可合理支持較低的取樣頻率。風險評估應考量取樣頻率降低亦會使更多批次產品暴露於風險中,且一旦發生問題,可能嚴重影響病患用藥供應。
Q12:Where should process gas be monitored?
製程氣體應於何處監測?
A: The monitoring of process gas should be performed as close as possible before the sterilization filter (the level of contamination before sterilization should be under control to ensure the efficiency of the gas sterilization process).
答:製程氣體的監測應盡可能於除菌過濾器前進行(除菌前的污染程度應受到管制,以確保氣體除菌製程的效率)。
4.4 Personnel/Training (Annex 1, Chapter 7)
人員/訓練(附錄 1,第 7 章)
Q13: This paragraph requires that all personnel accessing grade A and B areas be trained in aseptic gowning and aseptic behaviors. It also stipulates that compliance with the gowning procedure must be confirmed by means of assessments and periodic reassessments on an annual basis, covering both visual and microbiological checks (monitoring of gloved fingers, forearms, etc.).
本節要求所有進入 A 級和 B 級區域的人員,均須接受無菌更衣與無菌行為訓練。其亦規定,必須透過評估及每年定期再評估,確認是否遵守更衣程序;評估內容包括目視檢查與微生物檢查(如戴手套手指、前臂等之監測)。
Are these assessments to be covered by staff participation in APS? Paragraphs 9.38 and 9.39 mention staff participation in APS only in the context of staff requalification. Do APSs also have to take place during the initial qualification of employees? Does every employee have to perform every manual intervention in APS in order to be qualified or requalified?
這些評估是否應由員工參與 APS 來涵蓋?9.38 和 9.39僅在員工再資格確認的脈絡中提及員工參與 APS。APS 是否也必須於員工初始資格確認期間執行?是否每位員工都必須在 APS 中執行每一項人工介入,才能完成資格確認或再資格確認?
A: To ensure product quality, adequate training of employees working in grade B and A areas or involved in aseptic processes (incl. the necessary preparatory activities) is essential. The qualification must be adapted to the respective activities of the single employee and, after initial training (initial qualification), must also include regular requalification / retraining.
答:為確保產品品質,必須對在 B 級和 A 級區域工作,或參與無菌製程(包括必要準備活動)的員工提供適當訓練。人員驗證必須依員工各自執行的活動調整;在初始訓練(初次驗證)後,亦必須包括定期再驗證/再訓練。
Annex 1 requires that each employee qualified and involved in aseptic processes participates in a successful APS at least annually (or every six months if the aseptic processes are manual) as part of his requalification.
附錄 1 要求每位已完成資格確認且參與無菌製程的員工,至少每年參與一次成功的 APS(若無菌製程為手動操作,則每六個月一次),作為其再資格確認的一部分。
However, Annex 1 is not specific about the scope of the initial employee qualification, but indicates in paragraph 7.4 that the relevant training must cover theory and knowledge transmission as well as practical aspects and that evidence of training effectiveness is required through assessments using both visual and personnel monitoring examinations.
然而,附錄 1 並未具體規定員工初始資格確認的範圍,但於 7.4 中指出,相關訓練必須涵蓋理論、知識傳授及實務面向,並要求透過目視檢查與人員監測檢查等評估,提供訓練有效性的證據。
Although not explicitly required in Annex 1, the expectation for the initial qualification of an employee for the aseptic area is that practical process simulations, including manual interventions, are carried out under the supervision of qualified trainers/QA and are followed by personnel monitoring as training verification.
雖然附錄 1 未明確要求,但對無菌區域員工初始資格確認的預期,是在合格訓練人員/品質保證人員監督下,執行包括人工介入在內的實際製程模擬,並於其後進行人員監測,作為訓練確認。
It is at the discretion of the respective pharmaceutical company to define and justify whether these process simulations need to be conducted separately (but under similar conditions as an APS) or can be integrated within an APS. It is important that the representativeness of the activities to be performed by the trainee for the actual processes is justified and that the simulations cover each critical activity to be carried out by the respective employee. Equivalent representative interventions can be grouped for staff qualification. All operators should perform one intervention per year from each group of equivalent representative interventions.
各製藥公司可自行界定並說明這些製程模擬是否需單獨執行(但條件應與 APS 相近),或可整合於 APS 中。重要的是,必須說明受訓人員所執行活動對實際製程的代表性,且模擬應涵蓋該員工將執行的每項關鍵活動。可將等效且具代表性的介入操作分組,用於人員資格確認。所有操作人員每年應從每一組等效且具代表性的介入操作中,執行至少一項介入操作。